
Medical Equipment Manufacturers: Understanding Production Standards And Regulatory Compliance

Regulatory pathways, market access considerations, and post-market surveillance practices
Regulatory pathways for medical devices typically depend on device classification, intended use, and associated risk. Manufacturers may follow different conformity assessment routes such as self-declaration, third-party certification, or submission of clinical evidence, depending on regional rules and device risk categories. Early determination of the applicable classification and conformity procedures can help define the scope of required documentation, testing, and potential interactions with regulatory authorities. Manufacturers may also consider international regulatory requirements when planning market access in multiple jurisdictions.
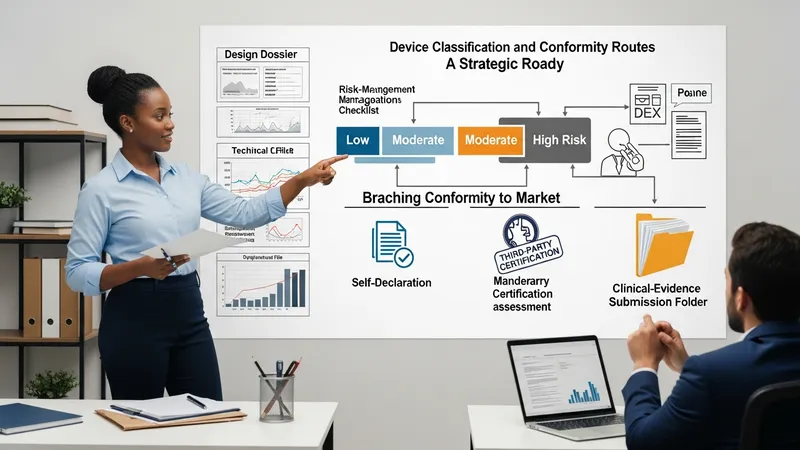
Market access planning often includes preparation of a technical file or design dossier that summarizes design rationale, verification and validation evidence, manufacturing controls, and risk management. For some devices, clinical data or literature may be part of the dossier to demonstrate performance under intended use. Manufacturers may maintain a regulatory affairs function to monitor changes in standards and regional regulations, since updates can influence labeling, post-market obligations, or required conformity assessments over time.
Post-market surveillance and vigilance processes are commonly required to continue assessing device performance after commercialization. Activities may include complaint intake procedures, adverse event reporting to competent authorities as applicable, periodic safety update reports, and trend analysis to detect changes in performance metrics. Corrective and preventive actions (CAPA) procedures provide structured methods to investigate root causes and document measures taken. These outputs can feed back into design, manufacturing, or labeling changes as part of continuous compliance management.
When devices are modified or new information emerges, manufacturers may reassess regulatory obligations and update technical documentation accordingly. Changes that affect safety or intended use may trigger additional testing, reclassification, or renewed conformity assessments. Manufacturers often document decision rationales and the associated evidence to demonstrate that regulatory requirements were considered during change implementation. Continued alignment of design, production, and post-market data supports sustained conformity across the product life cycle.